
 MARKING FOR MEDICAL DEVICES.
DELIVERED BY CE EXPERTS.
Expert conformity work for medical devices, from risk management to technical documentation. We prepare your assessment records, clinical or performance evaluation documentation and the declaration your device needs. Your dedicated expert coordinates the agreed work, giving your team clear evidence and a structured documentation handover.
Willy Lebherz
Founder and Managing Director of easyCE GmbH
DoC ready to sign!
We prepare your EU declaration of conformity and supporting documentation ready for the manufacturer to sign once the required assessment is complete.
Risk management documented!
We identify hazards, evaluate risks and document risk controls using ISO 14971, with traceability to requirements and verification.
Clinical evaluation prepared!
We prepare and review clinical evaluation documentation, including the clinical evaluation report, for your medical device.
UDI and registration organised!
We guide UDI assignment and prepare the information needed for the applicable EUDAMED registrations.
Quality management developed!
We develop your quality management system with reference to ISO 13485 and prepare the documentation needed for the applicable assessment route.
Trusted by leading businesses. Chosen for results.





Your role determines your medical device responsibilities
Medical device manufacturers
Medical device distributors
Medical device exporters
Medical device importers
Start with the intended purpose, classification and applicable regulation.
Medical devices fall under the Medical Devices Regulation (EU) 2017/745, while in vitro diagnostic medical devices are governed by Regulation (EU) 2017/746. The classification determines the conformity assessment route and whether a notified body must be involved.
Prepare the technical documentation, maintain risk and quality management processes, and establish the clinical or performance evidence appropriate to the device. Post-market surveillance, vigilance and the relevant registration and UDI duties continue after market entry.
easyCE prepares and reviews the agreed documentation, from risk management and safety requirements to instructions for use and the EU declaration of conformity. The manufacturer remains responsible for demonstrating conformity and signing the declaration.
Distributors must verify the required marking and information before supplying a device.
Check the CE marking, EU declaration of conformity and manufacturer information, together with importer details and UDI where required. Maintain the manufacturer's specified storage and transport conditions.
Keep records of complaints, non-conforming devices, recalls and withdrawals. If you suspect non-conformity, inform the relevant supply-chain partners and do not supply the device until the issue is resolved. Serious risks or suspected falsification also require notification to the competent authority.
easyCE reviews the documentation supplied with your devices and identifies the actions needed for your role. We help you organise evidence and respond clearly to compliance questions.
We match device identifiers across declarations, available certificates and accompanying information so your team can trace the evidence to the products it supplies.
Export requirements depend on the destination market.
CE marking indicates conformity with the applicable EU requirements. It does not replace the registrations, approvals or documentation that another country may require. Confirm the destination requirements and agree responsibilities with your customer or local partner.
A certificate of free sale may be requested for export. Under Article 60 of the MDR, the relevant Member State can issue this certificate at the request of an eligible manufacturer or authorised representative. It is not a universal requirement for every shipment.
easyCE prepares and reviews the agreed conformity documentation and helps you organise the product information needed for your export project.
We organise the agreed documents by device and destination, with clear version references for handover to your customer or local representative.
Importers bring devices from outside the EU onto the EU market and have their own verification duties.
Before placing a device on the market, verify the required conformity assessment, CE marking, declaration and product information. Check the manufacturer and its authorised representative, and confirm UDI and registration requirements where applicable.
Provide the required importer details, keep relevant declarations and certificates, and follow the manufacturer's storage and transport conditions. Handle complaints and suspected non-conformity, cooperate with the authorities and support corrective action when required.
easyCE checks your supplier documentation and identifies gaps before they delay market entry. We prepare the agreed technical records and review instructions for use against the applicable requirements.
Your CE project. Expertly handled.
Swipe or scroll horizontally to compare all options.
Approach to CE marking
Flexibility of support
Scope of support
Project contact
A clear, practical approach to CE marking
Our experts carry out the conformity work, with a clear plan tailored to your device
Choose the full agreed scope or individual assessment and documentation tasks
Your dedicated expert coordinates the agreed work throughout the project
Other CE service providers
Approach and project fit depend on the provider
Processes and flexibility depend on the provider
Scope agreed with the provider
Contact arrangements agreed for the project
CE software
A standardised workflow that your team adapts to the project
Your team manages the software and carries out the assessment
Features depend on the software and the modules purchased
Technical project support may require an additional service
We make CE marking simpler
Successful projects. Results that speak for themselves.
Conformity assessment for a dialysis water treatment system
easyCE supported the conformity assessment and prepared instructions for a permeate treatment system supplying water for dialysis equipment. The assessment considered the design measures and operating conditions needed to maintain water purity.
Technical requirements research for dental implants
easyCE researched technical requirements for a dental implant, including thread geometry, angles and material properties. The work addressed the information needed to assess design choices such as the use of titanium.
CE marking for horn spectacle frames and sunglasses
easyCE supported the conformity assessment of horn spectacle frames under the Medical Devices Regulation (EU) 2017/745 and sunglasses under the PPE Regulation (EU) 2016/425. We expanded the technical documentation, including the risk management and post-market surveillance plans.
Working with the customer, we reviewed the relevant general safety and performance requirements and the supporting design and manufacturing records.
We also assisted with the relevant registration processes, including EUDAMED actor registration for an SRN, the German DMIDS system and UDI assignment through IFA. These were separate tasks within the project.
Conformity documentation for a medical lubricant
In this historical project, easyCE assessed the requirements for a lubricant under the Medical Devices Directive 93/42/EEC and worked with the customer's existing quality management system. We prepared product-specific risk management records, hazard checklists, technical documentation and a declaration of conformity in German and English. The work also included documentation addressing the general safety and performance requirements in MDR Annex I.
Conformity assessment for a multi-drug test
easyCE reviewed the intended purpose of a multi-drug test and prepared the agreed conformity documentation. A test intended to examine human specimens for medical purposes is assessed under the IVDR; its classification depends on its intended use and the applicable rules. We prepared risk management and safety records, updated the technical documentation after design changes, coordinated with notified bodies and assisted with EUDAMED actor registration and UDI-related preparation.
CE marking for veterinary cauterisation devices
easyCE supported the conformity assessment of handheld veterinary cauterisation devices and prepared a declaration of conformity ready to sign. The project addressed electrical safety with reference to EN 60335-1 and electromagnetic compatibility, including shielding and interference suppression measures. EN 55014-1 was considered for emissions. Veterinary use does not itself bring a product within the human medical device regulations.
CE marking for an anaesthetic gas scavenging system
easyCE supported the conformity assessment and prepared a declaration ready to sign for an anaesthetic gas scavenging system. The assessment addressed the relevant safety and performance requirements in EN ISO 80601-2-13 and electrical safety with reference to EN 60601-1. The project included validation of gas extraction flow rates and documentation of the system's performance.
Your project in three steps
Step 1
Get free initial advice
Tell us about your product, your project stage and what you need help with. A CE expert will discuss the requirements and next steps with you, free of charge.
Step 2
Receive your proposal
Receive a proposal setting out the work your project needs and the scope of our service. We explain the details and answer your questions before you decide how to proceed.
Step 3
We get it done
Once you appoint us, your dedicated CE expert carries out the agreed work and keeps you informed throughout the process. Your expert remains available for follow-up questions after completion.
Medical device expertise. Delivered.
Frequently asked questions
Which EU regulations govern CE marking for medical devices?
The Medical Devices Regulation (EU) 2017/745 (MDR) applies to medical devices. In vitro diagnostic medical devices fall under Regulation (EU) 2017/746 (IVDR). Their general application dates were 26 May 2021 and 26 May 2022 respectively.
The regulations replaced the previous Medical Devices, Active Implantable Medical Devices and In Vitro Diagnostic Medical Devices Directives, subject to transitional provisions. Identify the intended purpose and classification before selecting the conformity assessment route.
Can devices assessed under the previous directives still be placed on the market?
Some legacy devices benefit from transitional arrangements, but not automatically. Eligibility depends on the device, its existing declaration or certificate, continued compliance, and the conditions and deadlines for moving to the new regulation. Certain MDR transitions run to the end of 2027 or 2028; IVDR transitions can run to the end of 2027, 2028 or 2029. Significant design or intended-purpose changes and missed conditions can affect eligibility. We review the specific position of your device.
Which medical devices require CE marking and a notified body?
Medical devices are generally subject to CE marking, with specific exceptions such as custom-made devices and devices used in clinical investigations. MDR risk classes are I, IIa, IIb and III; IVD classes are A, B, C and D.
For most class I medical devices, the manufacturer completes the conformity assessment without a notified body. A notified body is required for specified aspects of class I devices supplied sterile, with a measuring function, or as reusable surgical instruments, and for higher MDR classes. Under the IVDR, notified-body involvement is generally required except for non-sterile class A devices. The intended purpose and classification determine the route.
Do medical software and apps require CE marking?
Software requires CE marking under the MDR or IVDR when its intended purpose qualifies it as a medical device or an in vitro diagnostic medical device. Use in a hospital or a health-related app description alone does not establish this. We assess the intended purpose, functions and claims, identify the applicable classification and prepare the agreed conformity documentation.
Where should a medical device manufacturer start?
Start by defining the intended purpose, users and patient population, then establish the applicable regulation and classification. Identify the evidence needed for safety and performance, risk management, clinical or performance evaluation, technical documentation and quality management. easyCE turns these requirements into a clear project scope and carries out the agreed assessment and documentation work.
Expertise in product safety and compliance
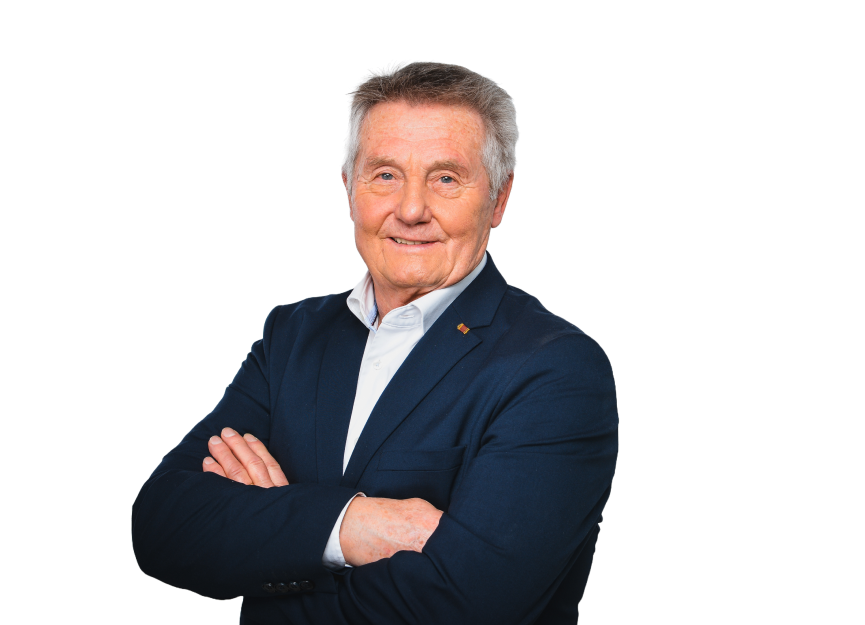
Willy Lebherz, Founder and Managing Director of easyCE
- Expert in product safety and compliance, since 1995
- Recipient of the "Medal of the Order of Merit of the Federal Republic of Germany", awarded in 1983 by the then Federal President Carl Carstens
- Meister qualification in measurement and control technology
- Captain (ret.), Project Officer for Technical Logistics in the Army Material Office and Chief of the Telecommunications Repair Company
easyCE specialises in product safety and conformity assessment. We combine engineering expertise with digital tools to help medical device manufacturers and their supply-chain partners bring compliant products to market. From risk assessment and identifying relevant standards to technical documentation and test coordination, we carry out the work your project needs. Choose the full agreed process or individual tasks, with a dedicated CE expert as your point of contact. Founded in southern Germany, we work with customers worldwide.
Tell us about your product. We will identify the requirements and the work needed to move your project forward.